2026 Stryker Knee Recall: How to Check Your Implant
Check the products covered by the 2026 Stryker knee implant recall, how to find your catalog and lot numbers, and what to do if they match.
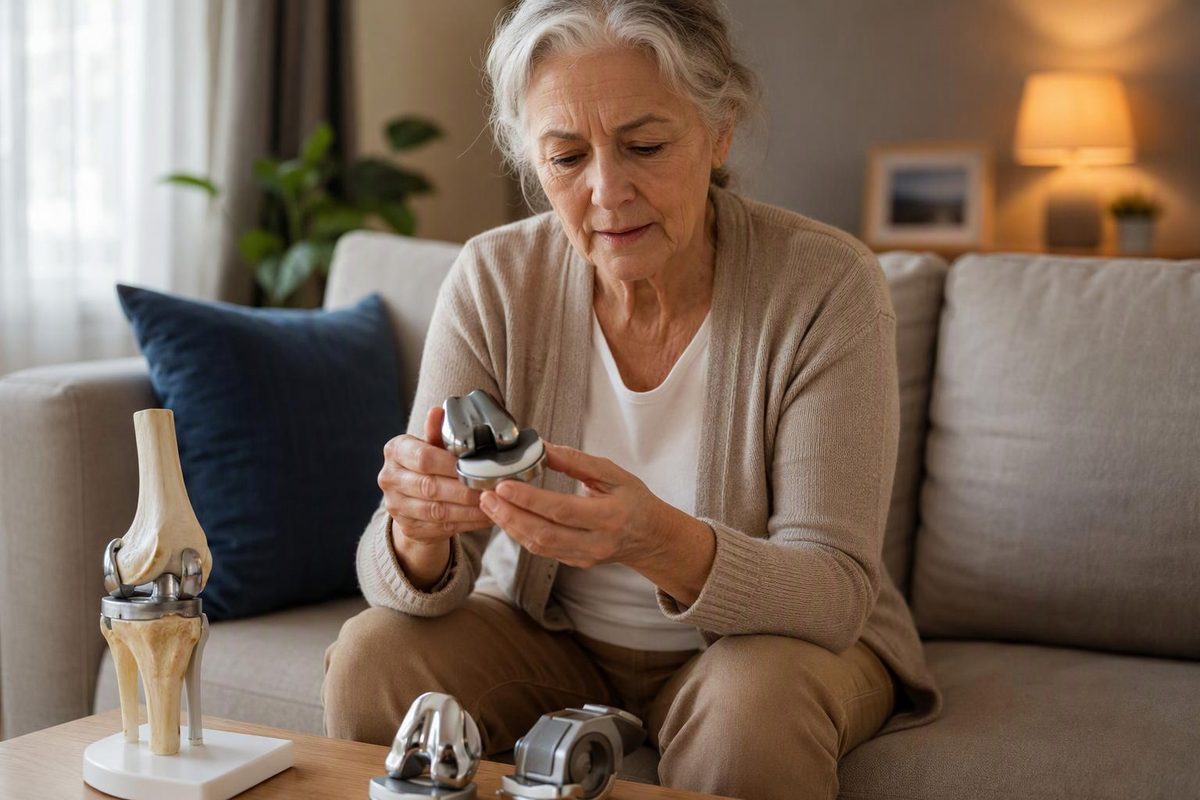
The 2026 Class II recall applies to specific catalog-and-lot combinations of Stryker knee components, not every Stryker knee replacement. Howmedica Osteonics initiated it on July 30, 2026, and the FDA posted it on September 16 as an open, classified recall covering 12 regulatory records across knee and other joint products (FDA event records).
Why the components were recalled
The affected components contain ultra-high-molecular-weight polyethylene (UHMWPE), a plastic used in joint-replacement bearing surfaces. According to the FDA, they were manufactured with UHMWPE raw material aged for more than five years.
The manufacturer reported that the material could have elevated oxidation, potentially increasing wear and leading to excessive wear debris, fragments, fracture or separation of a component. Possible harms listed in the recall are pain, inflammation and revision surgery (FDA recall Z-3197-2026). The record does not say that every affected component will deteriorate or fail.
This is a Class II recall. FDA defines that as a situation in which exposure may cause temporary or medically reversible harm, or in which the probability of serious harm is remote (FDA recall definitions).
Which Stryker knee products are included?
A product name alone cannot confirm that your implant is included. The catalog number and lot number must match an FDA entry.
The knee-related records include selected lots of:
- NRG Knee CR and NRG Knee P/S components
- Scorpio+ Mobile Bearing PCS
- Scorpio C-Dome, U-Dome, Inset, M-Dome and Recessed Patella components
- Scorpio Recessed Metal Backed Patella components
- Scorpio TS tibial inserts
- ScorpioFlex Total Knee CR components
- Series II tibial bearing inserts
- All Poly Constrained Insert 54 mm
- Triathlon Hinge Insert, size 4 in 11 mm and 13 mm
- selected specialty components bearing Duracon, GMRS, Kali, KRH, MRH and Triathlon names
This is a product-family summary, not a substitute for checking the identifiers. The FDA’s complete event search contains all 12 records and their catalog-and-lot combinations. More readable individual entries include the main NRG, Scorpio and ScorpioFlex group, Triathlon Hinge inserts and Scorpio metal-backed patella components.
The event also covers components for other joints. FDA displays “12,156 US” as the quantity in commerce on the recall records, but that is not a count of implanted knee replacements or affected patients. The public records do not state how many recalled components were implanted.
How to check your implant
- Find your implant card, if you received one. It may show the manufacturer and product identifiers, although it may not contain every number needed.
- Request the implant record from the hospital or surgeon. Ask for the implant log, implant stickers or operative report from your surgery. Specifically request catalog and lot numbers for the polyethylene tibial insert and any patellar component.
- Compare both identifiers with the FDA records. A matching product family with a different lot number does not establish that your component is recalled.
- Ask the surgical team to verify stock if your operation is upcoming. Howmedica told healthcare customers to quarantine affected products, discontinue their use and return them (FDA).
The FDA records name Howmedica Osteonics as the recalling firm and list 201-831-5000 for additional information. Your surgeon or hospital is generally the practical first contact because it holds the record of what was implanted.
Does a matching implant need to be removed?
Not automatically. FDA explains that an implanted device recall does not always require removal; the clinician and patient may need to compare the risk of leaving a device in place with the risks of another operation (FDA medical-device recall guidance).
The public action for this recall tells healthcare customers to return unused affected products. It does not direct every patient with a matching implanted component to undergo revision surgery, and it does not publish a single monitoring plan for all patients.
If your catalog and lot numbers match, contact your orthopedic surgeon even if the knee feels normal. Ask whether you need a clinical review, imaging or a changed follow-up schedule. Do not stop normal activity or arrange testing solely from the product name without first confirming the identifiers and discussing your circumstances with the surgical team.
When symptoms need attention
Contact your orthopedic team promptly if you develop a new or worsening change such as:
- increasing pain, swelling or warmth
- a new catching, shifting or unstable sensation
- declining walking ability after the knee had been stable
- a sudden loss of movement or function
These symptoms do not prove that a recalled component has oxidized or failed. Seek urgent assessment for sudden severe pain with inability to bear weight, an obvious deformity, or a hot red knee with fever; see the broader knee pain and swelling warning signs.
Patients and clinicians can voluntarily report suspected device problems through FDA MedWatch. A report does not by itself prove that a device caused an event, but reports contribute to post-market safety monitoring (FDA reporting guidance).